Contents
The pharmaceutical industry has seen what happens when qualification is skipped. There have been cases of contaminated medicine. Product was made on equipment that was never proved fit. Tests were not run before a drug was released. Today's rules come from failures like these. So a project is not finished until the client and the engineering team are both satisfied. Every piece of equipment has to be duly qualified. It has to comply with the prescribed principles and practices.
What equipment qualification is
Equipment qualification is the documented verification that a piece of equipment is fit for its intended use. The most important word here is "documented". An inspector cannot see what happened to a machine in the past. The inspector can only read its records. If a step was done but not written down, the inspector treats it as not done.
In short, qualification in the pharmaceutical industry means proving in writing that equipment is fit for its purpose. The proof is built stage by stage. Each stage is checked against a specification that was agreed before the equipment existed.
For this reason, qualification and validation in industrial pharmacy are taught as one subject. The equipment qualification definition on this page covers the half that comes first. The types of qualification in pharma are the stages of this proof. They run in a fixed order. Each stage is finished before the next one starts.
Equipment qualifications are written per machine. The plant does not share one file across its machines. A line of ten machines has ten files.
Equipment is qualified. That is different from being certified. Pharma equipment certification is a phrase that many people use. But no medicines regulator issues a certificate for a machine. A medicines regulator is the authority that must approve a drug.
Some certification schemes do exist, but for narrower purposes. None of them replaces qualification. A supplier can certify a material of construction or a calibration.
For pressure vessels, an Authorized Inspector stamps the vessel to the applicable section of the ASME Boiler and Pressure Vessel Code. The inspector also registers the vessel with the National Board. In the EU, some equipment needs a notified body's conformity assessment. A notified body is an organisation approved by an EU country to check products. This applies to higher-risk machinery categories and to higher-category pressure equipment under the Pressure Equipment Directive. Most process machinery is self-declared by the manufacturer.
None of these certificates proves that the specific installed machine does the job it was bought to do. Qualification proves that. It is what the medicines regulator looks for. So use these words with care. A loose word can be the difference between a file that holds up and an inspection finding.
Qualification and validation are not the same thing
Equipment is qualified. Processes, methods and cleaning are validated. Qualification proves that a thing is what it should be. Validation proves that an activity gives the same result every time. Qualification is one part of validation. This is why people often mix up the two words. It is also why the difference matters.
| Qualification | Validation | |
|---|---|---|
| Applies to | Equipment, instruments, utilities, facilities, computerised systems | Processes, cleaning, analytical methods, sterilisation, shipping |
| The question | Is this thing built, installed and operating as specified? | Does this activity give the same acceptable result every time? |
| Runs | URS, DQ, FAT, SAT, IQ, OQ, PQ | Protocol, consecutive runs, ongoing verification |
| Ends with | A qualified piece of equipment | A validated process |
| Depends on | Nothing before it | Qualified equipment. A process cannot be validated on unqualified equipment |
The last row matters most in practice. People use "equipment validation" to mean one of two things. It can mean the qualification described on this page. It can also mean the process validation that runs on the equipment afterwards. Neither can be done out of order. If a machine is not qualified, the process data from it proves nothing. The process being tested and the machine used to test it are both unproven.
Water systems are the clearest example of qualification and validation running together. They have their own sequence. Our guide to validating high-purity water systems explains the three-phase approach used to validate a PW or WFI loop. PW is purified water. WFI is water for injection. This validation builds on the equipment qualification described here.
The seven steps of equipment qualification
The equipment qualification flow chart runs URS, DQ, FAT, SAT, IQ, OQ and PQ, in that order. On a conventional project, each step must pass before the next one starts. A risk-based programme may combine stages that sit next to each other. For example, it is common to run SAT and IQ as one protocol. But the programme does not skip the evidence behind each stage. We follow these seven steps to make sure each piece of equipment performs reliably.
| Step | Where it happens | What it establishes |
|---|---|---|
| 1. URS, User Requirement Specification | Before anything is designed | What the equipment must do |
| 2. DQ, Design Qualification | Before anything is built | That the proposed design meets the URS |
| 3. FAT, Factory Acceptance Test | At the manufacturer's site | That it performs before it ships |
| 4. SAT, Site Acceptance Test | At the user site | That it still performs after delivery |
| 5. IQ, Installation Qualification | On site | That it is installed and configured as designed |
| 6. OQ, Operation Qualification | On site | That it operates across its specified range |
| 7. PQ, Performance Qualification | On site, with product | That it produces the required result in the real process |
1. Defining equipment expectation with User Requirement Specification (URS)
The URS is a formal document written at an early stage. It lists the general requirements and functional specifications for the equipment. It also lists what the organisation expects from the equipment. A well-written URS helps the supplier fully understand the client's needs. It also gives both parties a clear, short written record to work from.
The URS also helps in later stages of validation, such as installation qualification (IQ) and operational qualification (OQ). In the end, it helps build better quality products that suit the client's needs and meet the intended purpose. It also helps avoid disputes. Without a URS, the two parties may disagree or have conflicts of interest. With a URS, both parties agree on the same requirements.
The URS must include the objectives, technical specifications, design data, documentation, installation and commissioning, delivery, training, commercial terms and approvals. Equipment selection follows from the URS. Drafting the technical specs first stops a team from picking a machine first and then trying to justify it.
2. Verifying designs with Design Qualification (DQ)
After the User Requirement Specification is final, the design qualification is prepared. It is the main documented verification that the proposed equipment design meets the specifications and functions required. It also checks that the supplier meets regulatory requirements.
The DQ is finalised at this stage. The protocol formats for IQ, OQ and PQ are also drafted and agreed now. So the decision on the equipment, and on how it will be proven later, is made in the planning phase. This happens before any part of the machine is built. The project then runs against this whole DQ/IQ/OQ/PQ set. Any deviation in the equipment details is identified. Risks are assessed, documented and sent to the manufacturer.
Broadly, DQ has three steps:
- Checking that the supply meets the technical and financial terms in the purchase order.
- Verifying the specifications, reporting deviations, and sending them to the manufacturer.
- Writing a summary of the result.
DQ is also the stage where the pharmaceutical equipment design is fixed. After DQ, any change is a change to a machine that is already being built. This is why the URS must start from the process. An equipment catalogue is the wrong place to start. For example, first decide which of the granulation routes the product needs. If a granulator is specified before that decision, the machine is qualified against the wrong requirement.
3. Performance check with Factory Acceptance Test (FAT)
The factory acceptance test (FAT) is a functional test at the manufacturing site. It is done when the manufacturing of the machine is complete. The test fully checks the actual specification and functions of the equipment. It compares them with the datasheet, the specification and the purchase order.
FAT mainly inspects the quality and integrity of the equipment. It also reviews whether the equipment can run well in the conditions where it will operate later. All possible failures are looked at in advance. Modifications, adjustments or replacements are made as required. Fixing problems at this stage gives better control over the project, because the system is still with the manufacturer. The equipment has not yet arrived at the customer's site.
These points are kept in mind during a FAT:
- A detailed test plan. The plan sets out responsibilities, accountabilities and deliverables in detail. It helps measure performance. It also leaves room to find failures, issues or inefficiencies.
- Dry run testing. FAT is run empty (a dry run) or with substitute products. In exceptional cases, actual products may also be used for specific applications.
- Operators' involvement. Operators run the equipment. They must have the technical skills and know-how for the tasks. Involving and training them is essential. It makes sure that the equipment works the right way and is in the right hands.
- Leaving room for failures. The potential failures must be written down. A backup plan must be ready in case something unexpected happens. The plan stops the equipment from coming to a standstill.
- A detailed check for problems. After full testing, find the areas where the operator has difficulty and check them carefully. Answer how each problem will be fixed. Any gap in the system must be addressed so that the right answers can be found.
4. Verifying site functionality with Site Acceptance Test (SAT)
The Site Acceptance Test works on the same principle as FAT. Both tests check that the equipment functions correctly. The difference is where the test takes place. As the name says, SAT is done on site. The tests show whether the equipment received from the manufacturer meets its specifications.
The test includes a visual inspection of the components. It checks that the utilities work. It also checks that the equipment complies with overall safety and other usability parameters.
SAT is also the time to check that the operator can handle the equipment. Any further issues to be fixed are written down. In the end, the SAT gives both parties the final confirmation of performance. The tests mainly involve four steps. First, write a plan with the goals, objectives and approach of the SAT. Second, run the test cases. Third, write reports and document the results. Fourth, review the report and validate the results.
5. Checking the equipment setup with Installation Qualification (IQ)
Installation Qualification checks all aspects of the equipment. This documented verification checks the specifications, the design and the materials used, in detail. It also checks that delivery, installation and configuration follow the written, approved standards. Trained personnel carry out the installation. They make sure that the equipment and all its components have been delivered complete.
The tests check the following:
- Compliance with all specifications, and a review of design features for completeness and accuracy.
- Checking components for damage and reporting any damage.
- Gathering all manuals, designs and certifications.
- Setting up equipment maintenance, calibration schedules and procedures.
Re-qualification may be needed if a lapse or deviation is noted that may affect the quality of the equipment. In that case, a corrective action report is prepared.
6. Testing function reliability with Operation Qualification (OQ)
OQ starts after the IQ result has been reviewed and approved. OQ checks that the equipment can do the tasks it is meant for. The process involves:
- Testing the instruments.
- Finding parts that could malfunction.
- Checking that the main parameters and functions comply with the operating specifications.
- A thorough check, by the site representative, of the specification set during OQ.
- Reporting changes, if any, made to the system or equipment during OQ.
- Preparing and approving the Standard Operating Procedure (SOP) for operating and maintaining the equipment, instruments and systems.
Re-qualification is required if the equipment changes location or is moved. Modifications and newly added components also need to be qualified.
7. Checking real-world efficiency with Performance Qualification (PQ)
PQ is run after a successful SAT and an approved IQ and OQ. It measures the system's performance as defined in the protocol. It confirms that the equipment can operate in a specific environment. It tests how the real product performs under the equipment's real process parameters. The user prepares a Standard Operating Procedure (SOP) during the PQ.
The test shows that the system or equipment can make products with the specified quality and functionality, even in extreme cases. PQ can be seen as the final confirmation that the equipment is reliable. As with IQ and OQ, requalification may be needed if new equipment is added, because these changes can affect product performance. It may also be needed if a malfunction is found after PQ is done.
The guidelines it is written against
The seven steps are not our own convention. The GMP expectations of the major regulators have arrived at the same shape. GMP means good manufacturing practice. This is why the same sequence appears in a plant built for the US, the EU, the UK, Australia or a WHO-prequalified market.
- cGMP, in every jurisdiction that inspects the plant, requires premises and equipment to be suitable for their intended use. It also requires proof of that. cGMP means current good manufacturing practice. Equipment qualification is how that proof is given.
- The EU GMP guide has a separate annex on qualification and validation. Most European projects take their URS-to-PQ terms from it.
- USFDA, the US Food and Drug Administration, expects qualification as part of a lifecycle approach to process validation. In this approach, the equipment is qualified first. Then the process is shown to be reproducible on it.
- ICH quality guidelines set the wider framework, especially for risk management. ICH is the International Council for Harmonisation. Risk management decides how much testing a given piece of equipment needs.
- Client and contractual specifications also apply. The URS is a contractual document as well as a technical one.
One rule applies under all of them. The depth of qualification matches the risk. A tablet press touches the product. An air dryer three utilities away does not. The two do not get the same testing. The decision on how much to test is written down in advance, and it is part of the file.
Requalification, and what triggers it
Qualification is not done only once. Equipment qualified two years ago is qualified for the configuration, location and use it had two years ago. Requalification of equipment in pharma is the documented proof, done again after something changes.
These are the usual triggers:
- The equipment moves. A change of location, or a move within the same site, makes the installation evidence invalid.
- Something is modified or added. For example, a new component, a control system upgrade or a changed part that touches the product.
- The process changes. For example, a new product, a different batch size, or an operating range outside what OQ established.
- A deviation calls for it. A lapse is found during operation, and a corrective action report is attached.
- Periodic review. A review is scheduled at a frequency set by risk. It finds slow changes (drift) that nobody reported.
Requalification rarely repeats all seven steps. It repeats only the steps that the change affects. A change control is the formal written record of a change. A written change control that names the affected stages is more useful than a rule to redo everything.
Facility, utility and instrument qualification
The machine is not the only thing that must be qualified. Facilities, utilities and equipment qualification is the full scope. The same URS-to-PQ logic applies to each of them.
Facilities. Each room has a classification, a pressure cascade and a flow pattern. A pressure cascade means the air pressure steps down from cleaner rooms to less clean rooms. Each of these is qualified against the design before the first product runs. Facility qualification is what makes a finished room ready for use.
Utilities. Purified water, WFI, pure steam, compressed air and HVAC are each qualified as one complete system, with all its parts together. HVAC means heating, ventilation and air conditioning. Water systems are qualified and then validated over a long period. This is because their behaviour is seasonal and microbial, not mechanical.
Instruments. Instrument qualification, sometimes called analytical instrument qualification, is the same work on a smaller scale. A balance, a pH meter or an HPLC is qualified through design, installation, operational and performance stages. An HPLC is a lab instrument that separates and measures the parts of a sample. This qualification is in addition to routine calibration. Laboratory equipment qualification matters more than the size of the instruments suggests. A batch is released on the numbers these instruments give.
Computerised systems. Some machines have a control system that holds electronic records. In that case, the records themselves are also in scope. 21 CFR Part 11 is the expectation most projects are specified against. It is the US FDA rule for electronic records.
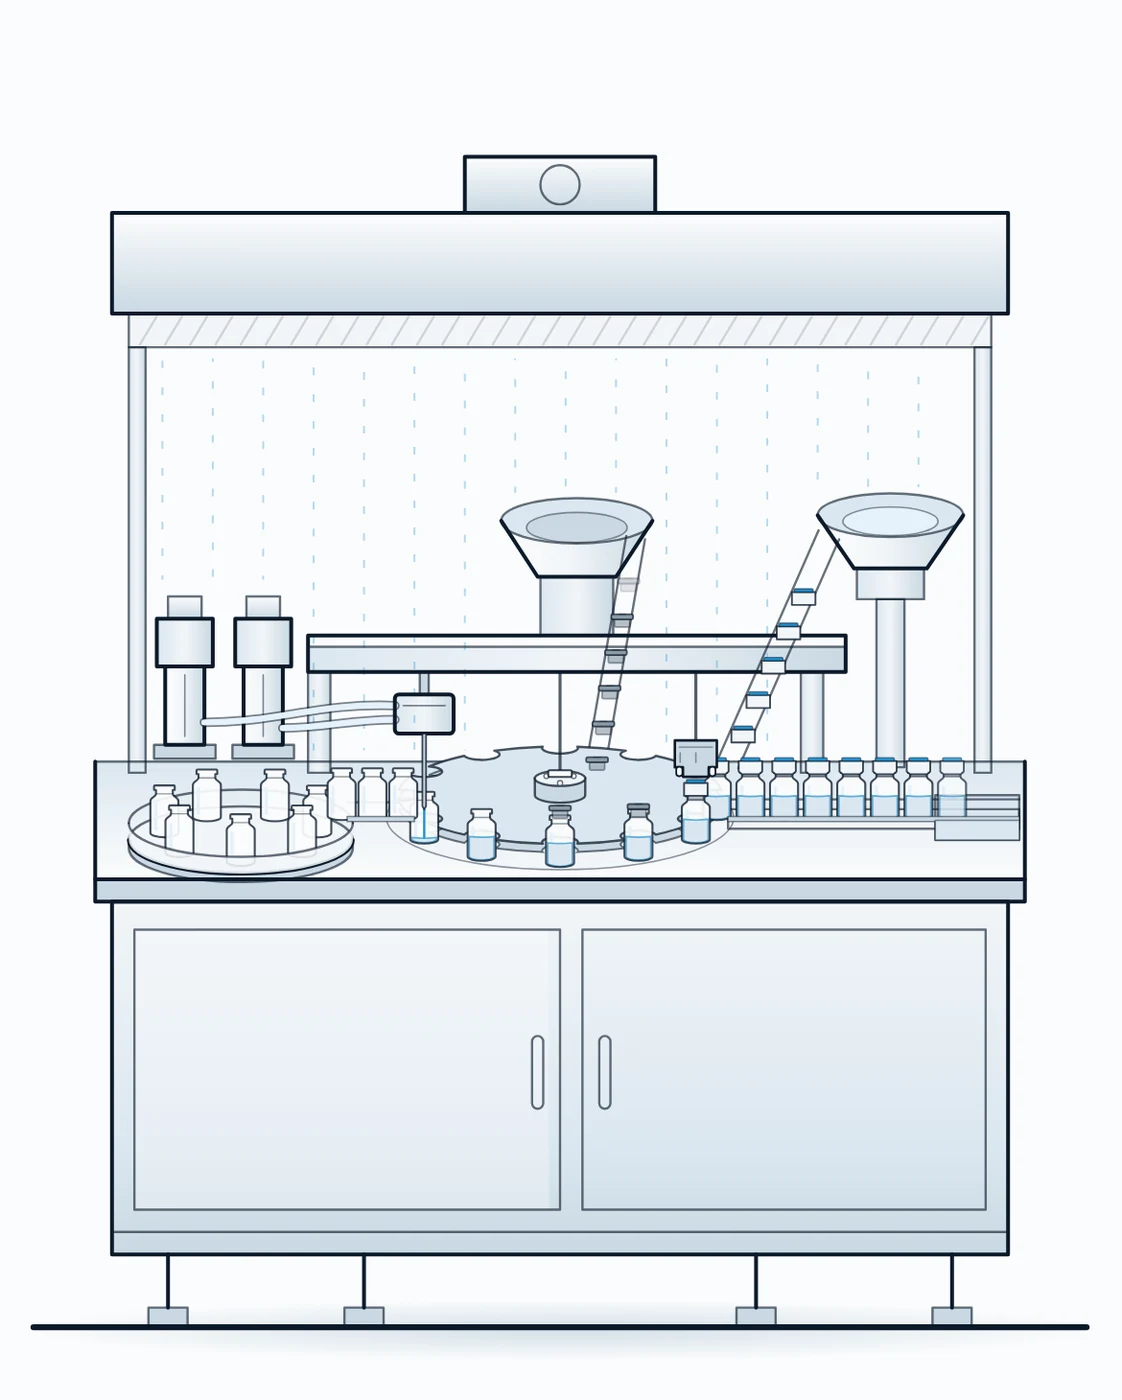
Aseptic filling equipment. A sterile line needs more work. The qualification must prove that the machine keeps the product sterile. It must also prove that the machine doses correctly.
This means media fills, which are test runs with a growth medium in place of the product. Where the machine runs in RABS (a restricted access barrier system) or an isolator, glove and barrier integrity is also tested. The performance qualification is carried out with the containers the line will run in production. Both the vial filling and capping machine and the prefilled syringe machine are handed over against this longer sequence.
Biotechnology and biopharmaceutical plants. Biopharmaceutical equipment qualification follows the same seven stages, but two things change. First, single-use assemblies move part of the evidence to the supplier. A single-use bag or filter arrives already certified and irradiated. It is not installed and tested on site like a fixed machine.
Second, biotechnology equipment qualification must also cover bioburden and endotoxin control. Bioburden is the number of microbes present. Endotoxins are toxic substances released from some bacteria. A small-molecule OSD (oral solid dosage) line, such as a tablet line, does not carry these controls. The sequence stays the same. The acceptance criteria change.
Who does what, and when
Qualification is shared work. Most problems in it happen at a handover, when work passes from one party to the next. Commissioning and qualification of pharmaceutical facilities is the industry's name for the combined activity. It is usually shortened to C&Q. Commissioning is the engineering work of making systems function. Qualification is the documented proof that they function as specified. On a well-planned project, the two run together.
| Who | Owns |
|---|---|
| The plant | The URS, the acceptance criteria, the final approval, and the SOPs that come out of OQ and PQ |
| The equipment manufacturer | The design against the URS, the FAT, the manuals, certifications and calibration records |
| The engineering partner | The protocols, the execution, the deviations, and the file that ties all of it together |
Fabtech runs this as part of building the plant.
Equipment qualification services and equipment validation services are usually bought separately. They come from a consultancy that arrives after the machines. When this work is done inside the project, the URS that starts the sequence is the same document the equipment was ordered against. The validation tests are part of Fabtech's own processes and protocols. They comply with GMP requirements for equipment validation in the pharmaceutical industry. They also comply with the user requirement specification and with contractual specifications.
We test the equipment so that it works as intended and meets its purpose at the customer's plant and beyond.
The equipment itself is on the page process equipment for pharma. When the scope is the whole plant, pharmaceutical turnkey projects covers the work from design to handover. The design stages that come before qualification are set out under pharmaceutical plant design. A line is handed over against the sequence on this page. For the wider rules, read good manufacturing practice guidelines for pharmaceuticals. Writing and running these protocols can be a full career. The page what to expect from a career in pharmaceutical engineering describes the validation and documentation career path and what employers ask for.
FAQ
What is equipment qualification in pharma?
It is the documented verification that a piece of equipment is fit for its intended use. It has seven stages: URS, DQ, FAT, SAT, IQ, OQ and PQ. It starts with a written specification, before anything is designed. It ends with a performance test on site, with real product.
What is the difference between qualification and validation?
Equipment, utilities, facilities and instruments are qualified. Processes, cleaning and analytical methods are validated. Qualification proves that a thing is what it should be. Validation proves that an activity reliably gives the result it should. A process cannot be validated on unqualified equipment.
What are the types of qualification in pharma?
The types are Design Qualification, Installation Qualification, Operation Qualification and Performance Qualification. The User Requirement Specification comes before them. The Factory and Site Acceptance Tests come between design and installation.
What are the seven steps of equipment qualification?
The seven steps are URS, DQ, FAT, SAT, IQ, OQ and PQ. Each step must pass before the next one starts. DQ is written against the URS. FAT is run against the design. SAT confirms that nothing changed in transit. IQ checks the installation. OQ checks the operating range. PQ proves the result with product.
What is installation qualification in the pharmaceutical industry?
It is the documented check that equipment was delivered complete and installed to the approved design. It verifies specifications and design features. It checks components for damage. It gathers manuals, designs and certifications. It also sets up maintenance and calibration schedules.
What triggers requalification of equipment?
Requalification is needed when the equipment is moved, modified or given a new component. It is also needed when the process or the operating range changes beyond what OQ established. A deviation with a corrective action attached, or a scheduled periodic review, also triggers it. Requalification usually repeats only the stages that the change affects.
Is pharmaceutical equipment certified?
No. Equipment is qualified, not certified. A supplier can certify a material or a calibration. An ASME Authorized Inspector stamps a pressure vessel. An EU notified body assesses higher-risk machinery and pressure equipment. But no medicines regulator issues a certificate for a machine. An inspector looks for a qualification file.
What guidelines is equipment qualification written against?
It is written against GMP in every jurisdiction that inspects the plant, and the EU GMP guide's qualification and validation annex. It also follows USFDA expectations for lifecycle process validation, and ICH quality and risk-management guidelines. The client's own URS and contractual specifications also apply.