Contents
What validation of a high purity water system is
Validation of a high purity water system is written proof that the system consistently makes water of the required quality. The proof covers the system as it was designed, installed and operated. It includes a design review and checks on installation and operation. It also includes a performance qualification, a long test that runs across all the seasons. Validation also covers sampling, maintenance and change control, which keep the proof up to date. Change control is the formal review of any change to the system before it is made.
The same method applies to purified water system validation and to pharma water system validation. It also applies to WFI system validation. WFI (water for injection) is the water grade used to make injectable medicines. Water validation in pharma differs from other industries in one way only: how much of it the inspector will read.
Why validation is mandatory
Regulatory guidelines around the world require every high purity water system to be validated. The type of system does not matter. Validation is needed to track how the system performs. This includes the storage and distribution systems.
Validation makes sure the system runs within its design parameters, the limits it was built for. It also makes sure the system consistently makes water of the desired quality. Validation also stops unacceptable microbial, chemical and physical impurities from forming. This applies while the water is made, stored and distributed.
Validation lets us prove how processes or systems will perform under all the conditions they will meet in future operation. The system shows this by consistently producing the same quantity and quality of water. The plant keeps it that way by following specific written operating and maintenance procedures. The high purity water systems market has grown with the biopharmaceutical industry. What regulators expect from validation has stayed the same for decades.
What validation of a high purity water system proves
Validation of high purity water systems involves proving:
- Engineering design
- Operating procedures and acceptable ranges for control parameters
- Maintenance procedures to accomplish it
The system has to be designed, installed and successfully tested in all the conditions it will run in. Changes in daily, weekly and annual usage patterns must also be validated.
A validation report is written at two points. The first is the end of equipment, instrumental and operational qualification. The second is when the validation protocol is complete. This report is the main document during an FDA inspection. It is also the reference document if the system is changed and revalidation (repeating the validation) is needed.
What the FDA Guide to Inspections of High Purity Water Systems looks for
Most inspectors follow the FDA Guide to Inspections of High Purity Water Systems, from the US FDA (United States). The guide tells an inspector what to look at:
- The system drawings, and whether they match what is installed
- The validation report and its data
- The microbial and chemical monitoring records, and how excursions (results outside the limits) were handled
- The sanitisation procedures and their frequency
- The maintenance and change records
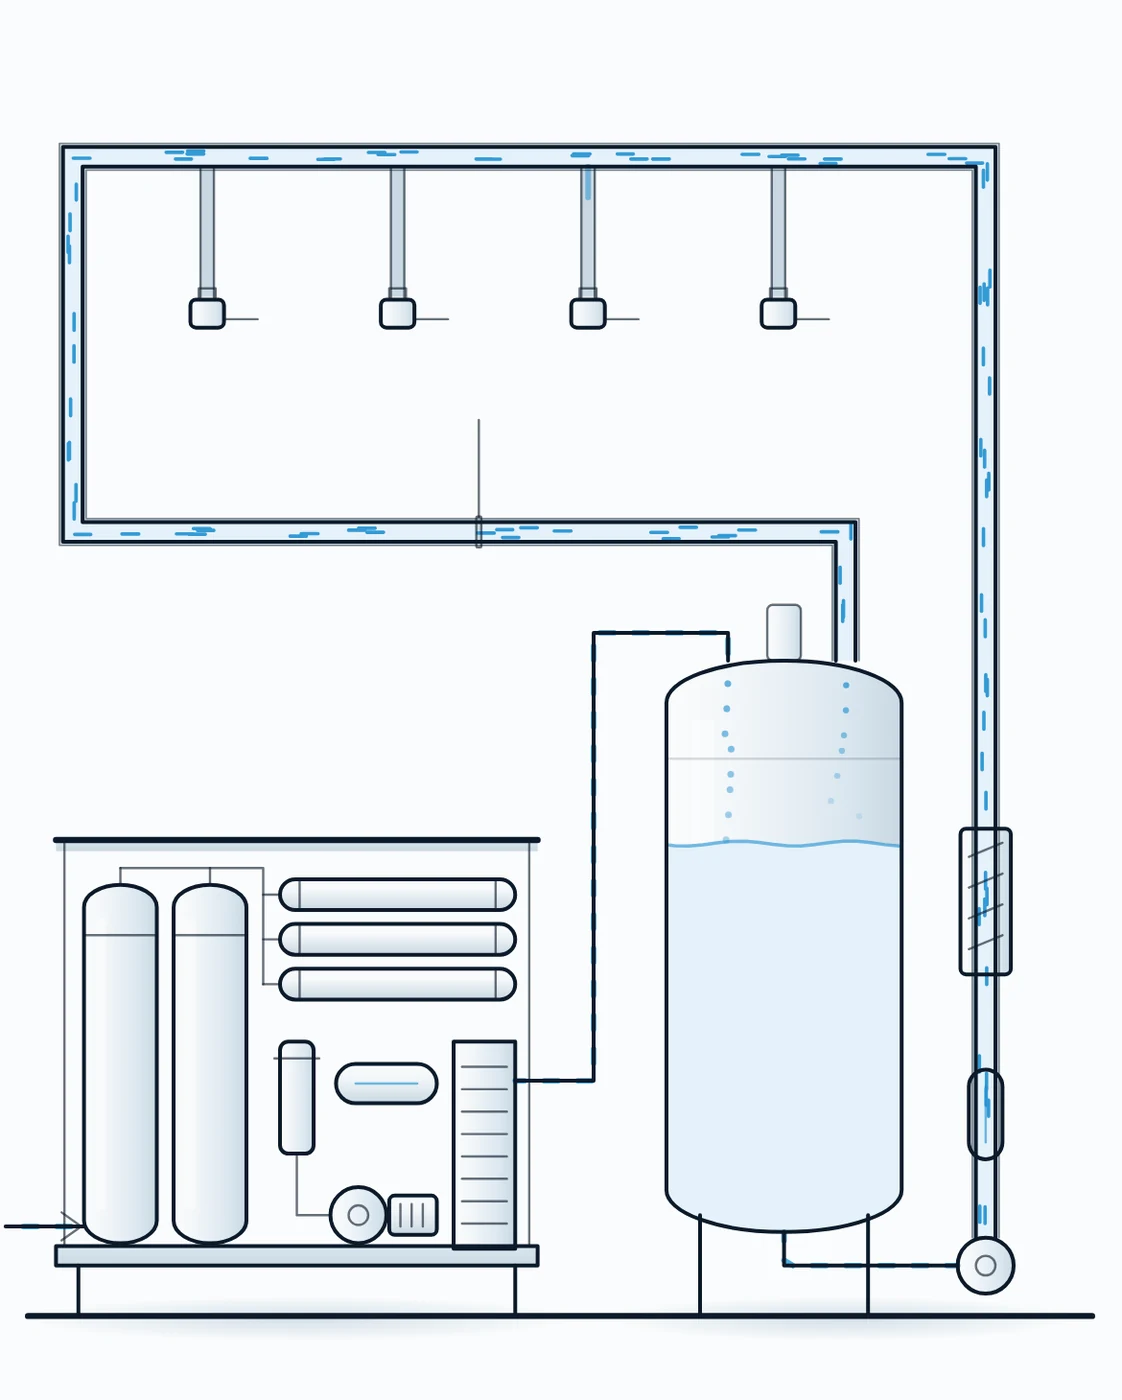
The guide looks closely at dead legs, loop velocity and how long water sits in tanks. A dead leg is a pipe branch where water stands still. Loop velocity is the speed of the water in the distribution loop. The guide also asks whether the alert and action limits came from real data. An alert limit gives an early warning. An action limit means the plant must act.
It is good practice to read the guide before the system is designed, not after. Most of the faults it finds are design faults. Sampling cannot fix a design fault later.
The three phases of performance qualification
The first two checks are installation qualification and operational qualification. They prove the system is built and works as designed. The performance check proves it keeps making the right water over time. It runs in three phases. The FDA guide and the ISPE Baseline Guide for water and steam systems both set out this approach.
| Phase | Duration | What happens | What it proves |
|---|---|---|---|
| Phase 1 | Two to four weeks | Intensive daily sampling at every point of use and after each treatment step; operating and sanitisation procedures finalised | The system can make the water and the procedures work |
| Phase 2 | Two to four weeks | Sampling continues at the same intensity while the plant runs the system under its final procedures | The procedures, not the commissioning team, produce the water |
| Phase 3 | About one year | Routine sampling at the frequency the plant will keep, through every season and raw-water change | The system holds its limits through the year; alert and action limits are set from real data |
Sampling plan: water validation in microbiology and chemistry
Water validation in microbiology proves that microbes in the water stay under control. It is the part most plants give too little time and effort. Chemical tests, conductivity and total organic carbon (TOC), can be read online. The control system records them and shows the trend. Microbial counts need a sample bottle, an incubator and days of waiting. So the sampling plan has to be made with care.
| What is sampled | Where | How often in Phase 1 and 2 | How often in Phase 3 and routine | Tests |
|---|---|---|---|---|
| Raw and pre-treated water | Inlet, after each pre-treatment step | Daily | Weekly to monthly | Chemistry, microbial count |
| Generated water | After RO, after EDI, after UV, after the still | Daily | Weekly | Conductivity, TOC, microbial count; endotoxin for WFI |
| Stored and distributed water | Tank, loop return, every point of use | Daily, each point of use | Points of use in rotation so each is sampled at least weekly | Conductivity, TOC, microbial count; endotoxin for WFI |
How a sample is taken matters as much as how often. Take each sample the way the water is used: from the same valve, after the same flush. Test it within the proven hold time. This is the time a sample can wait before testing without the result changing.
Documentation required: design, operation and validation
Documentation required during validation includes:
- Verification of design documentation including a description of installation and functions and specifications of functions
- Operating procedures
- Maintenance instructions & records
- Training of personnel
- Environmental records
- Plant inspection
Once validation is complete, the plant finalises an effective Standard Operating Procedure (SOP). The plant then has assurance that the system can consistently produce water that meets spec over long periods. If anyone sees a deviation from the SOP, the team must find out what changed and how it affects the whole system. The team then carries out revalidation based on that change and its impact.
So pharmaceutical water system design, operation and validation form one file, not three. The design documents say what was meant to happen. The operating records say what did happen. The validation report links the two.
Validation for MHRA, USFDA and other regulators
The rules are the same under every regulator. The wording and the focus differ. USFDA (United States) inspectors work from the guide above and from the plant's own data.
MHRA is the medicines regulator of the United Kingdom. MHRA-regulated UK manufacturing sites follow UK GMP (good manufacturing practice) and the European Pharmacopoeia water monographs. A monograph is the official quality standard for one grade of water. UK GMP stays closely aligned with the EU GMP annexes through PIC/S.
MHRA inspectors tend to focus on data integrity, which means records that are complete and unchanged. They also tend to ask about the reasons behind the alert and action limits. And they ask what the site does when a result is out of range.
High purity water system qualification follows the same three-phase approach here. Validation support for MHRA-regulated UK sites uses it as well, with the report written to the EU GMP structure.
WHO (World Health Organization) and TGA (Australia) inspections follow the same water system validation guidelines in their main points.
Commonly overlooked considerations
Any validation protocol must include the elements above. But validation takes a long time, and small details are often missed. A couple of the points most often missed are described below.
UV lights
UV (ultraviolet) light passes through microorganisms (microbes). It disrupts the DNA of the cell and inhibits reproduction. So the UV lamp must work at its best efficiency at all times. This keeps the UV output at the right level and keeps microbial contamination under control. The team takes water samples from test ports before and after the UV unit. The lab tests them to find out whether microbial contamination is present.
UV systems never remove all microbes. They reduce them by 2 to 3 logs. The reduction is sometimes around the level of 100 cfu/ml (colony-forming units per millilitre).
Standard validation procedures say UV lamps must be replaced on a fixed schedule, typically every six months to a year. This is because the UV output of a lamp falls over its rated life. The quartz sleeve is the clear tube around the lamp. It collects impurities from the raw water. The team cleans and inspects it separately, because scale on the sleeve also cuts UV transmittance (how much UV light passes through).
The ballast, which supplies power to the UV lamp, should also be changed every 2 to 3 years. Intensity meters measure the UV output, and they are a must. They help with the maintenance of the UV lamp and also with the validation.
Biofilm formation
Biofilm is a well-known problem in the water industry. It forms if water is stagnant (stands still), even for a short while. Bacteria, microbes, pathogens (E-coli, salmonella) and other organisms in the water attach to the inner surface of the pipelines. There they form biofilms. A new biofilm starts as a slimy layer. These layers are hydrophobic (they push water away), and they help the microbes stay attached to the surface.
Biofilm also damages pipes. It contains bacteria that produce hydrogen sulphide. The hydrogen sulphide reacts with iron in the pipes and causes corrosion. Removing biofilm is expensive and difficult, so prevention is always the preferred route.
Cleaning and sanitising the RO system
RO (reverse osmosis) is the membrane step that removes dissolved salts. To control microbial growth, the RO system must be cleaned and sanitised regularly, particularly 1st pass RO. Biofilm protects the bacteria and stops the sanitising reagent from reaching them. Cleaning before sanitisation helps break up this biofilm. Cleaning also removes foulants, the deposits that react with the sanitising reagent and use it up chemically.
Sanitising is a two-step process. First, an acidic reagent such as citric acid removes inorganic foulants. Then a high pH reagent such as sodium hydroxide helps remove organic foulants.
The RO system has traditionally been sanitised with formaldehyde or hydrogen peroxide. Modern polyamide membranes are usually sanitised with hydrogen peroxide or peracetic acid blends instead. The team then rinses the system well until all chemicals are flushed out.
This chemical cleaning and sanitisation must be carried out at regular intervals. The schedule should be based on the microbial levels in samples from the 2nd pass RO permeate outlet. Permeate is the clean water that has passed through the membrane.
Typically, under a standard SOP, cleaning and sanitisation follow a 10 to 15 day cycle once trend data is established. The plant's own data sets the real interval. The team should keep a detailed log, with the periodic test results that show any increase in microbial growth in the permeate water.
Annual revalidation: who does it and what it covers
Who does annual validation for high-purity fluid systems? Usually the plant's own quality and engineering teams do it. The system's maker supports the yearly requalification of the equipment. This covers instrument calibration, proof that sanitisation still works, and filter and lamp changes. It also covers a review of a year of data against the alert and action limits.
Annual validation for high-purity fluid systems is less work than the original three phases, but it is not optional. If anyone changes the system during the year, a change-control assessment comes first.
Selecting a packaged system that will validate cleanly
A packaged system is supplied as one assembled unit on a skid (a frame). What features should you prioritise when selecting a packaged high-purity water system for biopharmaceutical process validation? Choose the features that make the proof easy to show. The table lists them.
| Feature to prioritise | Why it matters at validation |
|---|---|
| Sampling valves at every treatment step and every point of use | Without them the sampling plan cannot be executed |
| Online conductivity and TOC with data logging | Continuous, attributable chemistry records instead of manual logs |
| A loop with no dead legs and a designed return velocity | The biofilm failure mode is prevented by design, not by sanitisation alone |
| Hot water sanitisable RO and loop | Sanitisation efficacy is proven by time and temperature, which are easy to record |
| Weld, boroscopy and material records delivered with the skid | The installation qualification is largely pre-written |
| A control system with audit trail and user access levels | Data integrity questions are answered before they are asked |
Monitoring and validating lab water dispensing systems
Lab water is often forgotten because the volumes are small. How to monitor and validate a lab water dispensing system follows the same method as a plant system, at laboratory scale. Define the water grade the tests need. Sample the dispensing point on a schedule. Trend conductivity and microbial counts over time. Record every cartridge, filter and lamp change. A QC (quality control) lab that cannot show this for its own water will find it hard to defend its test results.
Why validation matters
The quality of the high purity water is critical to making pharmaceuticals. Two things are essential to keep the quality and integrity of the water. The first is validation of the high purity water system. The second is close adherence to the SOPs. When you carry out a high purity water system validation, the points in this article will increase the chance of success.
This guide applies to the systems on our purified water generation systems and WFI generation systems pages. The beginner's guide to purified water generation systems explains how they work. The full range is on solutions for water.
FAQ
What is validation of a pharmaceutical water system?
It is written proof that the system consistently makes water of the required quality, as designed, installed and operated. It covers design review, installation and operational qualification, and a performance qualification run across the seasons. It also covers the sampling and maintenance that keep the proof up to date.
What does the FDA Guide to Inspections of High Purity Water Systems cover?
It lists what an investigator examines. This includes system drawings against the installation, the validation report and data, and monitoring records and excursion handling. It also includes sanitisation procedures and frequency, and maintenance and change records. The guide also covers design points such as dead legs, loop velocity and how long water stays in tanks.
How long does performance qualification of a water system take?
It takes about a year. Two intensive phases of two to four weeks each show that the system and its procedures work. A third phase of routine sampling then runs through every season. This lets the plant set the alert and action limits from real data.
Who does annual validation for high-purity fluid systems?
Usually the plant's own quality and engineering teams do it. The equipment manufacturer supports requalification. This covers calibration, sanitisation efficacy, filter and lamp changes, and a review of the year's monitoring data against the alert and action limits. The review looks for drift, seasonal excursions and any point of use that trends worse than the loop return.
What is water validation in microbiology?
It is the part of validation that proves microbial control. Every treatment step and point of use is sampled on a set schedule. The samples are cultured and the counts are trended. The record shows sanitisation keeps them below the action limit. It is slower than chemical testing, and plants most often give it too few resources.
What features should I prioritise when selecting a packaged high-purity water system for validation?
Prioritise sampling valves at every step and point of use, and online conductivity and TOC with logging. Also choose a loop without dead legs, hot water sanitisation, complete weld and material records, and a control system with an audit trail. Each one makes a piece of the proof easier to produce.
How is a water system validated for MHRA-regulated UK sites?
Support for MHRA-regulated UK sites can follow the same three-phase performance qualification. The report is written to UK GMP, which is aligned with EU GMP, and to the European Pharmacopoeia monographs. MHRA inspectors focus on data integrity, the reasons behind alert and action limits, and how excursions are investigated.
How do you monitor and validate a lab water dispensing system?
Define the water grade the tests need. Sample the dispensing point on a schedule. Trend conductivity and microbial counts, and record every cartridge, filter and lamp change. The method is the same as for a plant system, at laboratory scale. The dispensing point is treated as the single point of use that has to prove its water quality.